YM-58483 (BTP2) BTP2 Calcium Channel inhibitor suppresses astrocytic store-operated calcium entry and neuroinflammation
Copy code to add this image to your page
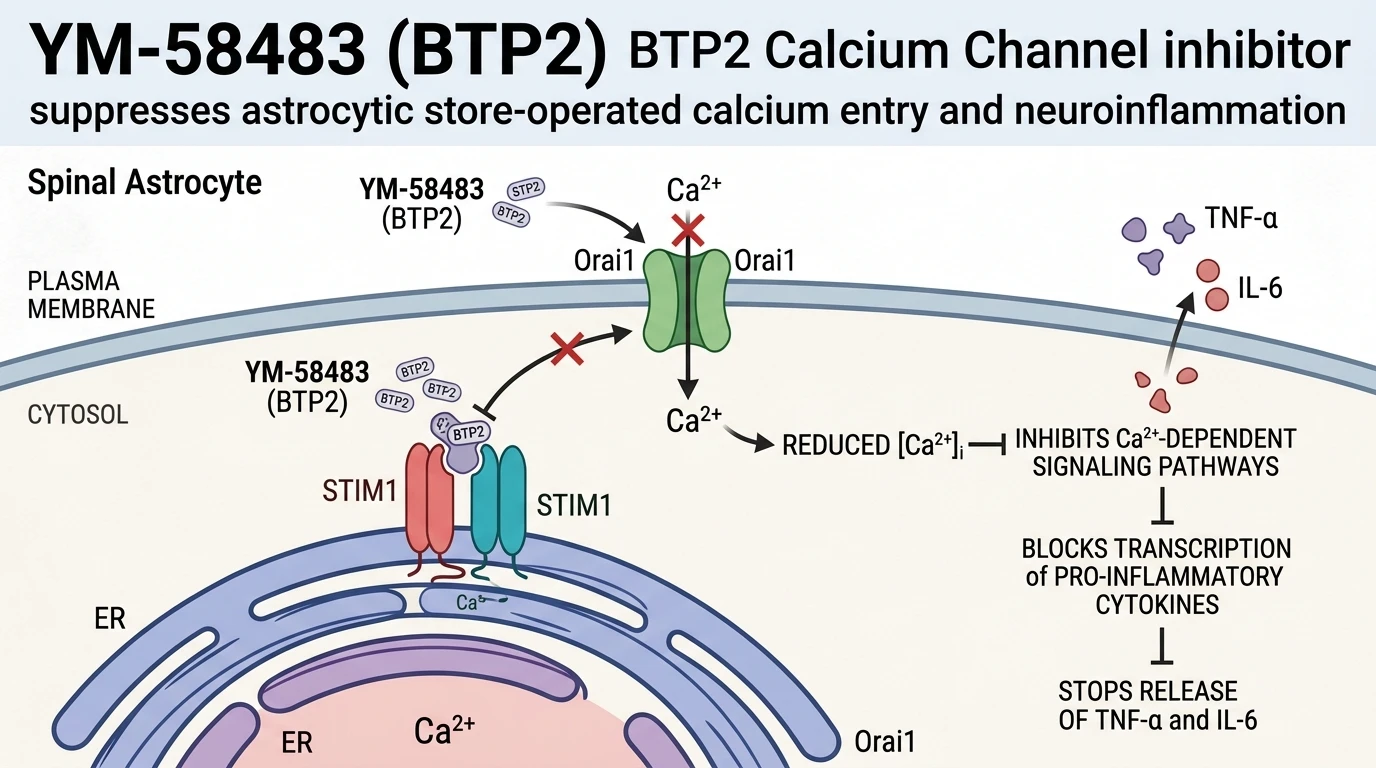
Primary spinal astrocytes utilize store-operated calcium entry (SOCE) to transduce luminal endoplasmic reticulum stress into cytosolic signaling cascades. Depletion of intracellular calcium stores prompts STIM1 and STIM2 sensors to oligomerize. These proteins redistribute toward junctional regions abutting the plasma membrane. Subcellular rearrangement leads to direct physical coupling with Orai1 pore subunits. The assembly forms functional calcium release-activated calcium (CRAC) channels. Extracellular calcium rapidly enters through Orai1 pore complexes. This influx drives pro-inflammatory gene transcription.
Small-molecule targeting of store-operated channels presents a defined strategy for modulating glial activation. The compound YM-58483 (BTP2) operates as a selective inhibitor of CRAC channel conductance. As a recognized BTP2 Calcium Channel inhibitor, this pyrazole derivative blocks store-operated Ca2+ entry following luminal store depletion. Experimental evaluations in cultured rodent spinal astrocytes confirm high potency under micromolar concentrations. Calcium imaging assays demonstrate complete suppression of SOCE amplitude following exposure to thapsigargin or cyclopiazonic acid.
Spinal astrogliosis contributes directly to central sensitization and chronic pain signaling. Astrocytic release of tumor necrosis factor-alpha (TNF-α) and interleukin-6 (IL-6) enhances synaptic excitability in dorsal horn neurons. Sustained store-operated calcium entry serves as the primary driver for cytokine transcription in these glia. Pharmacological blockade with YM-58483 (BTP2) disrupts this pathobiological sequence. Treatment with the compound suppresses inflammatory mediator synthesis without compromising basic cell viability. Comparisons with established chemical probes, such as small-molecule kinase inhibitors used in broader signaling studies, demonstrate distinct selectivity for ion pore entry over broad enzymatic inhibition.
Pharmacodynamic studies in isolated astrocyte populations highlight precise concentration-dependent profiles. Luminal depletion mediated by sarco/endoplasmic reticulum Ca2+-ATPase (SERCA) inhibitors induces steady calcium accumulation. Pre-incubation with YM-58483 (BTP2) prevents this rise across varied inflammatory stimuli, including lipopolysaccharide exposure. Quantitative real-time polymerase chain reaction and enzyme-linked immunosorbent assays confirm concomitant reductions in TNF-α and IL-6 expression. The block occurs without altering baseline resting calcium concentrations or mitochondrial membrane potentials.
Molecular target validation confirms STIM1, STIM2, and Orai1 expression in spinal astrocytic membranes. Short hairpin RNA knockdown of STIM1 significantly diminishes store-operated influx. Similarly, dominant-negative Orai1 mutants reproduce the pharmacologic phenotype observed with YM-58483 (BTP2). These findings establish that physical coupling between STIM sensors and Orai pores governs astrocytic calcium influx. Inhibiting this interaction attenuates downstream transcription factor activation, specifically nuclear factor of activated T-cells (NFAT).
Methodological assessments verify that YM-58483 (BTP2) efficacy remains consistent across diverse induction protocols. Whether store depletion occurs via chemical SERCA inhibition or receptor-mediated activation, the compound exerts uniform channel blockade. MTT cytotoxicity assays demonstrate preserved cellular viability across all tested pharmacological ranges. These observations support CRAC channel targeting as a focused intervention for glial inflammatory pathways.
Pharmacological Characterization and Neuroinflammatory Cytokine Suppression
Copy code to add this image to your page
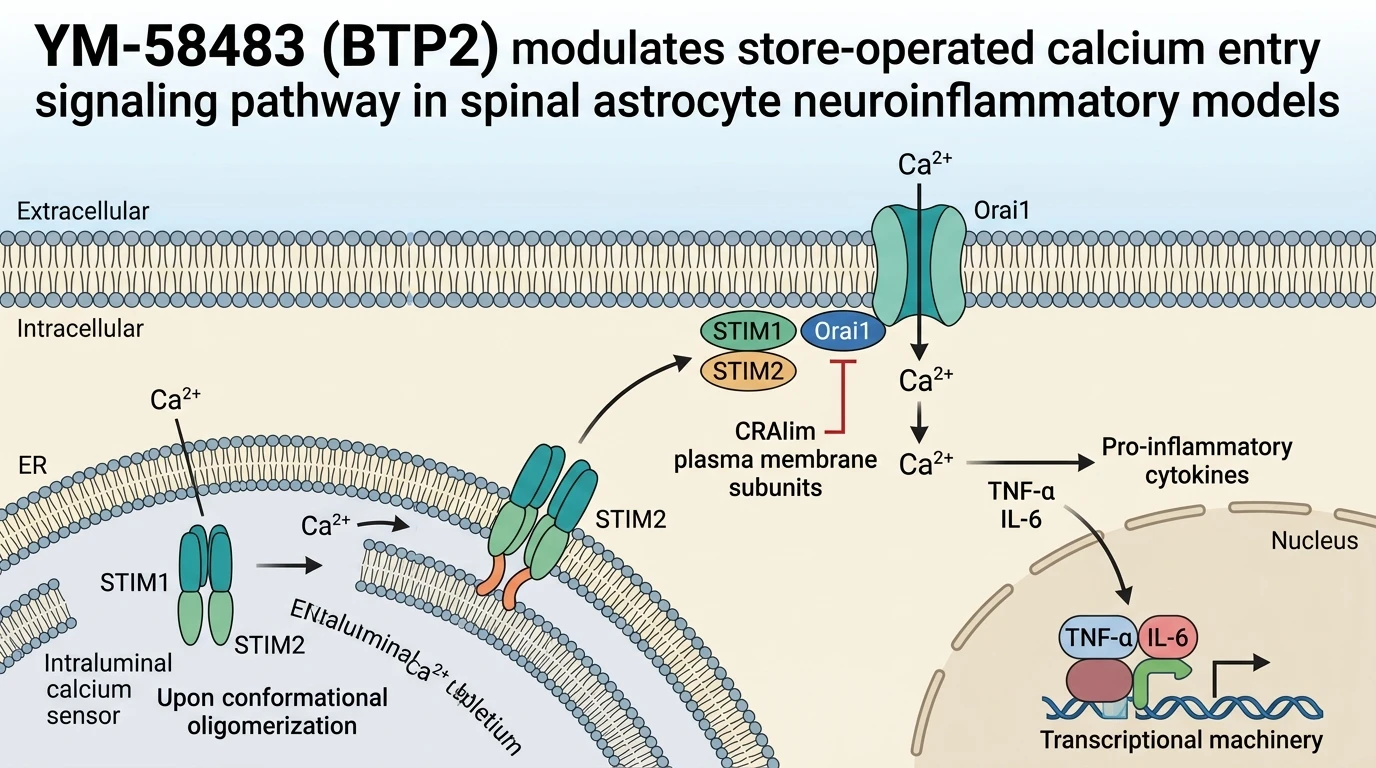
YM-58483 (BTP2) is a potent inhibitor of store-operated calcium channels (SOCCs) and CRAC channels formed by STIM1, STIM2, and Orai1 subunits. By blocking store-operated Ca2+ entry (SOCE) upon endoplasmic reticulum calcium depletion, YM-58483 attenuates downstream signaling cascades in spinal astrocytes. Pharmacological inhibition suppresses the transcription and secretion of key pro-inflammatory cytokines, specifically TNF-α and IL-6, induced by thapsigargin, cyclopiazonic acid, or lipopolysaccharide, without impairing cell viability.
STIM1 and Orai1 Coupling Mediates Astrocytic Calcium Entry
Luminal ER calcium depletion induces rapid STIM1 aggregation in spinal astrocytes. Translocation to puncta coordinates direct binding with membrane-bound Orai1 subunits. Fluorescence ratio imaging confirms robust intracellular calcium elevation following store depletion. YM-58483 (BTP2) application blocks this influx completely within minute timeframes. The BTP2 Calcium Channel inhibitor acts directly on CRAC channel complexes without altering baseline resting calcium levels.
Attenuated Cytokine Expression Following Chemical Store Depletion
Astrocytic cytokine release drives secondary dorsal horn hyperexcitabilities. Thapsigargin and cyclopiazonic acid stimulate pronounced upregulation of TNF-α and IL-6 mRNA. Pre-treatment with YM-58483 (BTP2) reduces cytokine protein secretion in primary astrocyte cultures. The compound achieves half-maximal inhibition at sub-micromolar concentrations under in vitro testing protocols. Cytokine reduction mirrors the kinetic profile of channel inhibition.
Preservation of Spinal Astrocyte Viability During Channel Inhibition
Sustained channel inhibition must not induce non-specific neurotoxicity. Cell viability assays confirm high survival rates following prolonged compound exposure. Lactate dehydrogenase release remains baseline across effective therapeutic dose ranges. The BTP2 Calcium Channel inhibitor preserves structural astrocyte integrity during inflammatory lipopolysaccharide challenge. Target engagement remains highly selective for SOCE mechanisms.