Ro-3306 CDK inhibitor Reverses Paclitaxel-Induced Mitotic Catastrophe and Apoptosis in Carcinoma
Copy code to add this image to your page
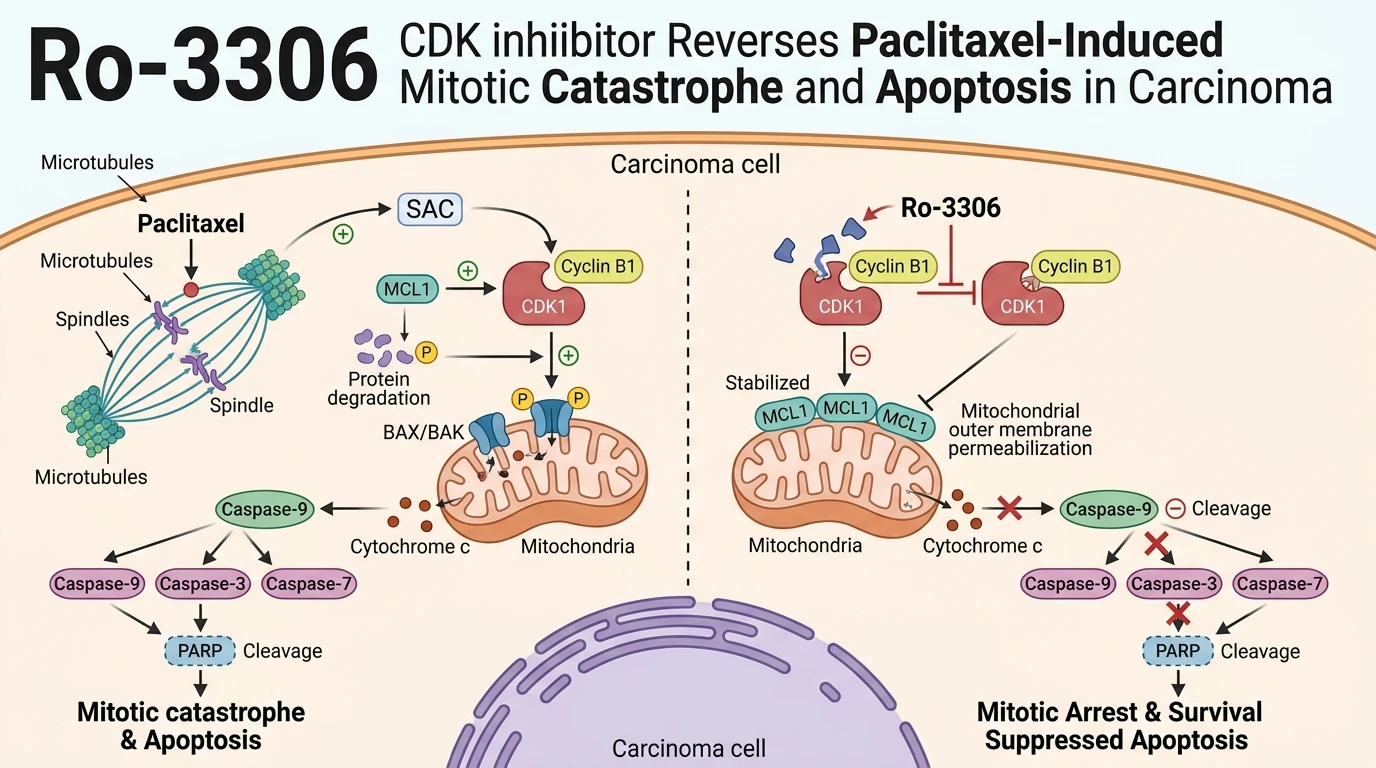
Cyclin-dependent kinase 1 (CDK1) functions as the primary enzymatic driver of the eukaryotic G2/M transition. Partnered with its regulatory co-factor cyclin B1, active CDK1 phosphorylates structural and regulatory substrates required for nuclear envelope breakdown, centrosome separation, and mitotic spindle assembly. Microtubule-stabilizing chemotherapeutics such as paclitaxel induce prolonged mitotic arrest by activating the spindle assembly checkpoint (SAC). Sustained SAC activation forces cells into either apoptotic death or premature exit from mitosis—a phenomenon termed mitotic slippage. However, the precise biochemical requirements for paclitaxel-driven cytotoxicity depend heavily on persistent CDK1 kinase activity.
The small-molecule ATP-competitive inhibitor Ro-3306 serves as a selective agent for dissecting CDK1-dependent biochemical signaling. As a targeted Ro-3306 CDK inhibitor, this compound demonstrates high selectivity for CDK1 over related cyclin-dependent kinases such as CDK2 and CDK4 under in vitro enzymatic assay conditions. Pharmacological assays in nasopharyngeal carcinoma (NPC) cell models show that co-administration or pre-treatment with Ro-3306 fundamentally alters cellular responses to microtubule toxins. Rather than enhancing tubulin-targeted cytotoxicity, Ro-3306 attenuates paclitaxel-induced lethality.
This pharmacodynamic antagonism stems from the absolute requirement of CDK1 activity for paclitaxel-triggered apoptotic cascades. When NPC cells encounter paclitaxel alone, tubulin polymerization is hyper-stabilized, preventing proper kinetochore-microtubule attachment. Continuous activation of CDK1/cyclin B1 complexes triggers phosphorylation events that signal downstream proteolytic machinery. Specifically, active CDK1 promotes the degradation of myeloid cell leukemia 1 (MCL1), an anti-apoptotic protein belonging to the BCL-2 family. Depletion of MCL1 compromises mitochondrial outer membrane integrity, leading to cytochrome c release, apoptosome assembly, and sequential activation of caspase-9 and caspase-3. Cleavage of poly(ADP-ribose) polymerase (PARP) follows, executing irreversible cell death.
By blocking CDK1 catalytic activity, Ro-3306 interrupts this apoptotic cascade at its apex. At physiologically relevant concentrations in vitro, Ro-3306 prevents paclitaxel-induced MCL1 degradation. Sustained levels of MCL1 preserve mitochondrial membrane potential and suppress pro-apoptotic pore formation. Consequently, cleavage of caspase-9 and PARP is markedly reduced. Cellular viability assays reveal that cells treated with the combination display reduced mitotic catastrophe and decreased overall cell death compared to those treated with paclitaxel alone.
These observations highlight the complex role of CDK1 in chemotherapeutic response. While CDK1 hyperactivation drives cell proliferation, its activity during spindle toxin arrest acts as a necessary executioner of cell death. Pharmacological blockade using the Ro-3306 CDK inhibitor shifts cellular fate from apoptotic clearance toward G2 arrest. Broader inquiries into nuclear machinery and structural dynamics, such as those conducted in cellular biology and genomic regulation research, emphasize the context-dependent nature of kinase inhibition.
Furthermore, this antagonistic relationship underscores critical considerations for combination drug regimens. Administering CDK1 inhibitors alongside spindle assembly toxins like paclitaxel may inadvertently protect malignant cells from chemotherapeutic lethality. In nasopharyngeal carcinoma models, active CDK1/cyclin B1 signaling is mandatory for paclitaxel-mediated killing. Consequently, temporal orchestration or strict sequential dosing strategies must be evaluated if CDK inhibitors are to be combined with microtubule-targeted agents.
Pharmacological Evaluation of CDK1 Inhibition and Antagonism
Copy code to add this image to your page
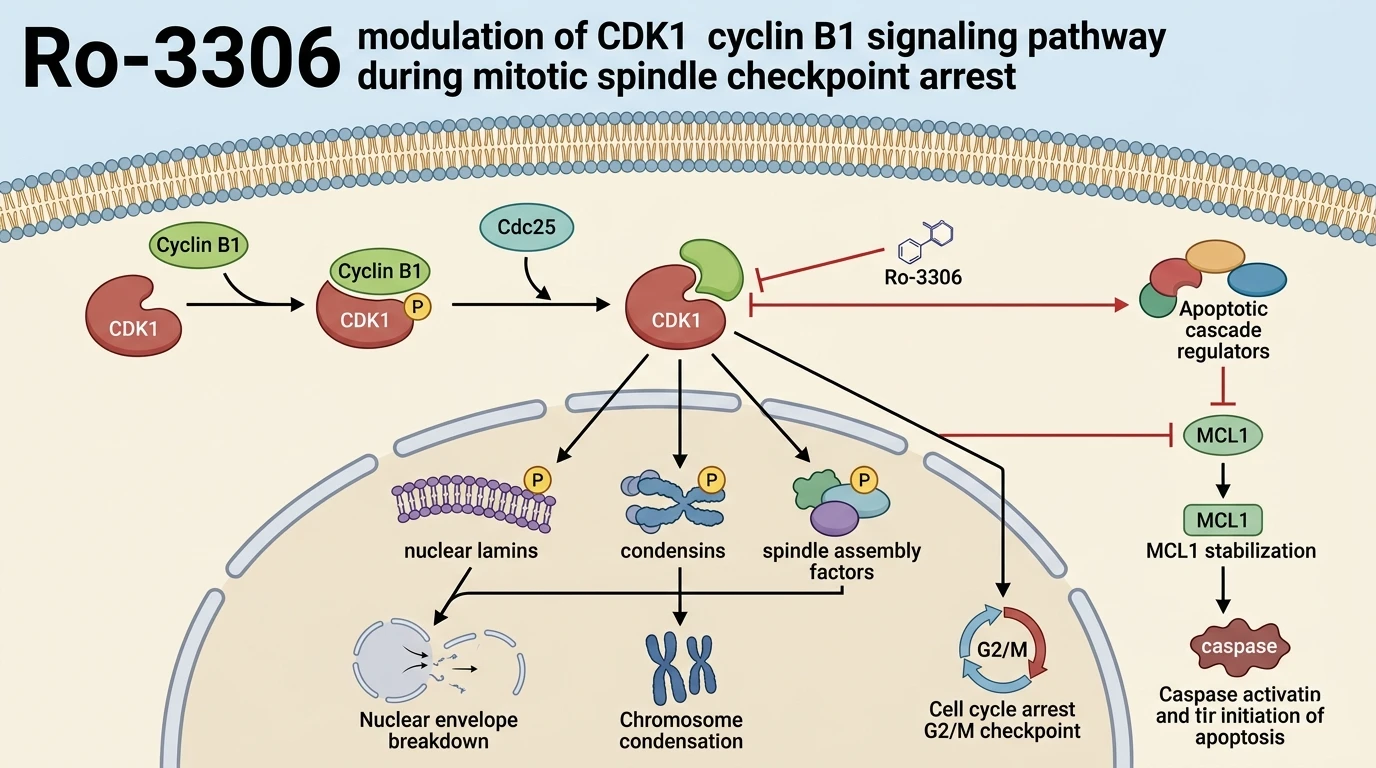
Ro-3306 selectively inhibits CDK1 kinase activity, disrupting CDK1/cyclin B1 signaling required for mitotic progression. In nasopharyngeal carcinoma models, Ro-3306 prevents paclitaxel-induced apoptosis by suppressing MCL1 proteolysis and blocking downstream cleavage of caspase-9 and PARP. This targeted blockade eliminates mitotic catastrophe, resulting in functional antagonism when combined with taxane-based chemotherapeutics.
MCL1 Protein Preservation and Caspase-9 Blockade
Inhibition of CDK1 catalytic activity by the Ro-3306 CDK inhibitor directly prevents the proteolytic clearance of MCL1 in paclitaxel-challenged nasopharyngeal carcinoma cells. Under standard paclitaxel treatment, active CDK1 phosphorylates priming sites that signal MCL1 for ubiquitin-mediated proteasomal destruction. Pharmacological suppression by Ro-3306 maintains MCL1 protein levels. Mitochondrial outer membrane integrity remains preserved. As a result, pro-apoptotic signaling stalls prior to apoptosome recruitment. Immunoblotting analysis demonstrates a pronounced reduction in cleavage fragments of both caspase-9 and PARP. This biochemical blockade effectively halts the apoptotic execution program in vitro.
Antagonistic Co-Administration in Nasopharyngeal Carcinoma
Simultaneous administration of Ro-3306 and paclitaxel yields an antagonistic survival outcome in experimental cell culture models. Paclitaxel relies on sustained CDK1/cyclin B1 activation to trigger mitotic catastrophe. When Ro-3306 suppresses CDK1 kinase activity, cells bypass microtubule toxin-induced mitotic death. Instead, cells accumulate in G2 phase without entering catastrophic mitosis. Cell proliferation assays confirm enhanced survival relative to paclitaxel monotherapy. These findings demonstrate that active CDK1 signaling serves as an essential mediator of taxane lethality in nasopharyngeal carcinoma cells.