Decitabine from Selleck Inhibits DNMTs to Restore Tumor-Suppressive miR-146a Expression
Copy code to add this image to your page
Aberrant DNA hypermethylation executed by DNA methyltransferases (DNMTs) represents a primary epigenetic mechanism governing tumor suppressor gene silencing during prostate cancer progression. Epigenetic alterations often precede genetic mutations, establishing transcriptional silencing at specific regulatory loci required for cellular differentiation, apoptosis, and androgen receptor signaling control. In advanced disease, particularly castration-resistant prostate cancer (CRPC), transcriptional suppression of microRNA species accelerates metastatic dissemination and treatment resistance. Among these non-coding transcripts, microRNA-146a (miR-146a) functions as a key negative regulator of pro-survival pathways by modulating inflammatory and oncogenic transcription factor networks.
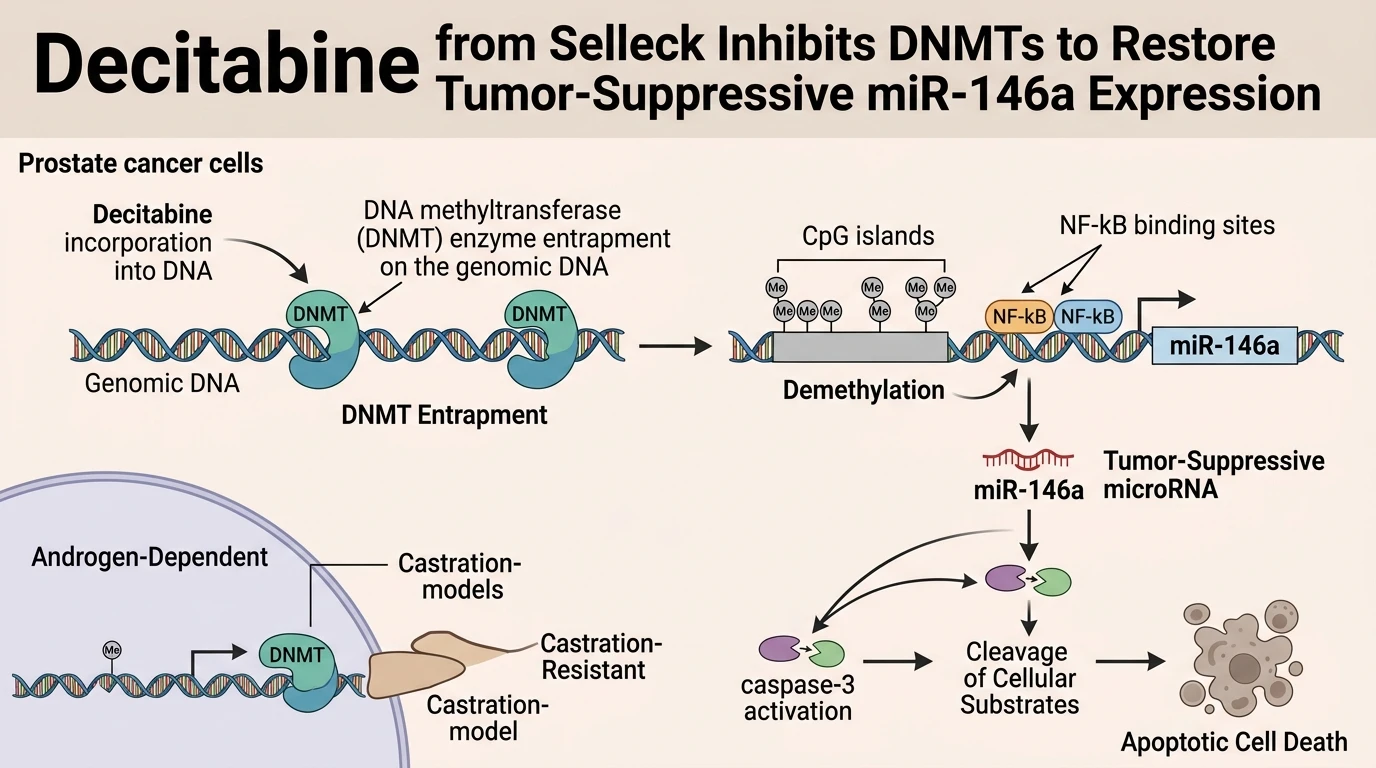
Decitabine (5-aza-2'-deoxycytidine) operates as a nucleoside analog inhibitor of DNMTs. Upon intracellular uptake via human equilibrative nucleoside transporters (ENTs), Decitabine undergoes sequential phosphorylation by deoxycytidine kinase to yield its active triphosphate form, 5-aza-2'-deoxycytidine triphosphate. During the S-phase of the cell cycle, this nucleotide analog substitutes for natural deoxycytidine residues within nascent genomic DNA. When catalytic DNMT enzymes (DNMT1, DNMT3A, and DNMT3B) engage the azacytosine ring during DNA replication, the modified cytosine ring forms a stable, covalent adduct with the active-site cysteine residue of the enzyme. This entrapment prevents the beta-elimination step required for catalytic release, physically locking DNMTs onto the DNA strand.
The formation of covalent DNMT-DNA adducts triggers rapid proteasomal degradation of the entrapped enzymes. Consequently, target cells suffer severe, dose-dependent depletion of soluble DNMT pools. As DNA replication continues across successive cellular divisions, maintenance methylation fails, causing a passive, replication-dependent reduction in genomic 5-methylcytosine levels across hypermethylated promoter regions. Utilizing high-grade reagents such as Decitabine from Selleck ensures precise target engagement without structural batch variability in cell-based assays.
Recent mechanistic evaluations demonstrate that Decitabine directly reverses hypermethylation at the miR-146a promoter locus in human prostate carcinoma models. Under basal conditions, high methylation density within CpG islands upstream of the miR-146a transcription start site suppresses its synthesis. Application of Decitabine downregulates promoter methylation, relieving transcriptional repression. Restored expression of mature miR-146a directly downregulates key target molecules within the NF-kB axis and associated signaling pathways, disrupting anti-apoptotic cellular programs.
In experimental assays spanning both androgen-dependent (LNCaP) and androgen-independent (PC3) human prostate cancer cell lines, exposure to Decitabine restores physiological levels of miR-146a transcript. This transcriptional reactivation precipitates a cascade of apoptotic events characterized by proteolytic cleavage of procaspase-3 into active caspase-3 fragments. Cleavage of downstream substrates, including poly(ADP-ribose) polymerase (PARP), leads to irreversible commitment to programmed cell death. Crucially, this biological response occurs in both hormone-sensitive and androgen-insensitive backgrounds, indicating that DNMT inhibition circumvents classical androgen receptor resistance pathways.
In vivo studies utilizing castrated nude mouse xenograft models bearing LNCaP tumors further corroborate these in vitro observations, as supported by recent epigenetic prostate cancer therapeutic evaluations. Monotherapy with surgical castration often results in transient regression followed by tumor recurrence driven by CRPC cells. However, combining surgical castration with systemic Decitabine administration suppresses tumor growth and prevents secondary tumor regeneration. Biomarker analysis reveals a pronounced reduction in serum free prostate-specific antigen (FPSA) levels in combination-treated cohorts relative to monotherapy controls. Bisulfite sequencing of genomic DNA isolated from xenograft tissues demonstrates sustained hypomethylation across critical CpG sites adjacent to the NF-kB binding element within the miR-146a promoter.
These pharmacodynamic findings establish that targeted DNMT inhibition by Decitabine restores endogenous miR-146a expression through localized promoter demethylation. By targeting regulatory CpG islands controlling microRNA transcription, Decitabine re-sensitizes refractory prostate cancer cells to apoptotic stimuli, offering an epigenetic approach to delay the transition from hormone-sensitive disease to castration-resistant states.
Pharmacodynamic Characterization and In Vivo Antitumor Efficacy
Copy code to add this image to your page
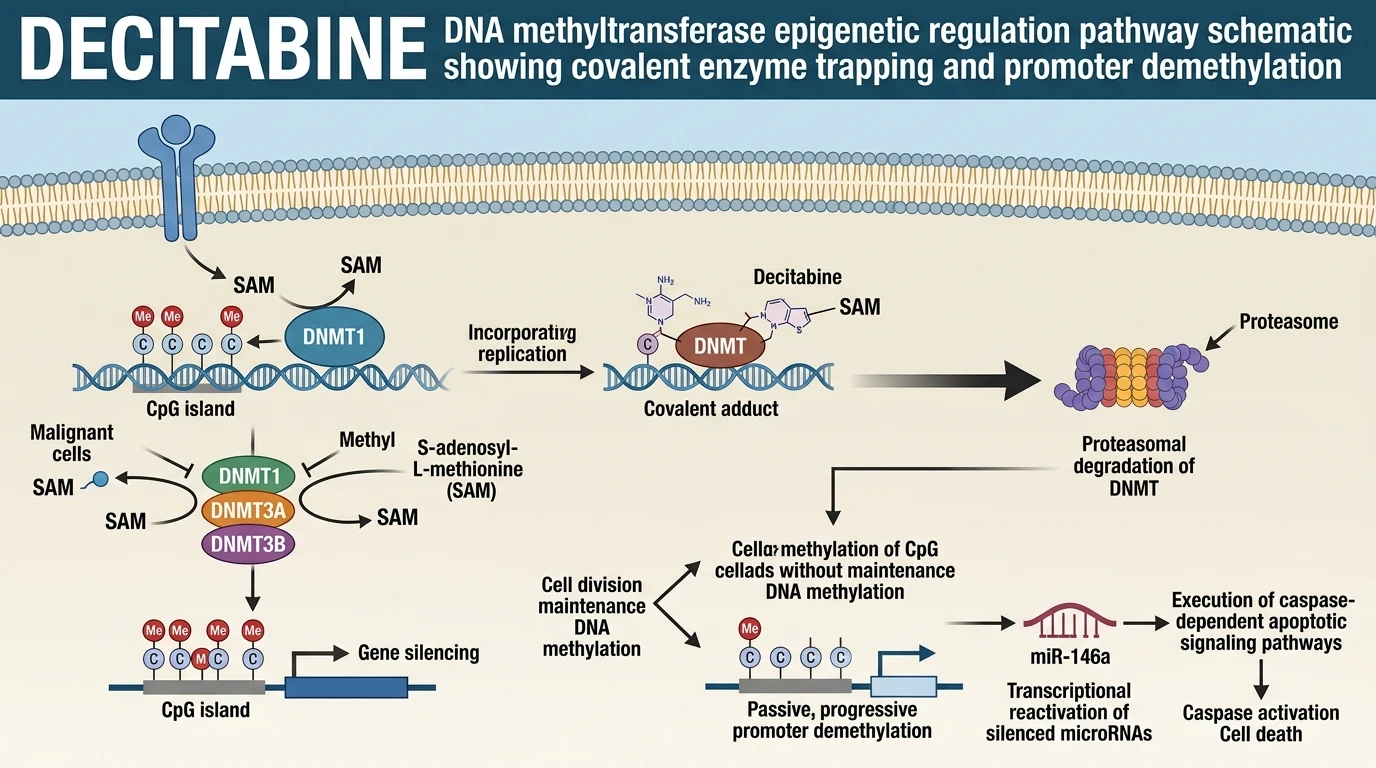
Decitabine inhibits DNA methyltransferases (DNMTs) by incorporating into DNA as a nucleoside analog and trapping catalytic enzymes in covalent DNA-protein complexes. Proteasomal degradation of trapped DNMTs causes passive, replication-dependent promoter demethylation. In prostate cancer models, Decitabine targets hypermethylated CpG sites adjacent to the NF-kB binding locus within the miR-146a promoter. Selective demethylation upregulates tumor-suppressive miR-146a expression, driving caspase-3 activation and inducing apoptosis in both hormone-sensitive and castration-resistant cell phenotypes while suppressing tumor growth and lowering serum free prostate-specific antigen levels in vivo.
Promoter Demethylation and Caspase Activation in Prostate Lines
In vitro administration of Decitabine effectively depletes functional DNMT enzymes in both LNCaP and PC3 human prostate cancer cell models. Methylation-specific PCR analysis confirmed a marked loss of 5-methylcytosine density across CpG islands regulating the miR-146a transcription start site. Quantitative reverse transcription PCR revealed a corresponding four- to six-fold increase in mature miR-146a levels following 48 hours of exposure under tested in vitro assay conditions. Downstream signaling assays demonstrated immediate proteolytic processing of procaspase-3. Caspase-3 cleavage was observed within 24 hours of compound treatment. Enzymatic fluorometric assays showed heightened caspase activity; this elevated activity preceded nuclear fragmentation and phosphatidylserine externalization. Importantly, cell viability dropped sharply in both androgen-dependent and androgen-independent phenotypes. These observations indicate that miR-146a promoter hypomethylation initiates cell death independently of functional androgen receptor signaling pathways.
Xenograft Growth Suppression and Marker Reduction in Vivo
In castrated nude mice bearing established LNCaP xenografts, combined administration of castration and Decitabine from Selleck delivered substantial therapeutic suppression. Combined androgen deprivation and methyltransferase inhibition halted tumor growth. Tumor volume remained static throughout the 28-day treatment period. Vehicle-treated castrated controls exhibited rapid tumor regrowth within 12 days post-castration. Serum analysis confirmed a drop in free prostate-specific antigen (FPSA) levels in combination-treated animals. Bisulfite genomic sequencing of excised tumor tissue identified consistent demethylation at key CpG loci adjacent to the promoter NF-kB binding site. The selectivity of demethylation was confined to hypermethylated CpG clusters without inducing global genomic hypomethylation toxicity. These combined data demonstrate that targeted epigenetic editing via Decitabine from Selleck prevents CRPC emergence within this murine xenograft model.